Run copy-number alteration inference
Usage
RunCNV(
  object,
  method = c("copykat", "fastCNV", "scevan", "infercnv", "numbat"),
  assay = NULL,
  layer = "counts",
  group.by = NULL,
  reference.by = NULL,
  reference = NULL,
  genome = c("hg38", "hg19", "mm10"),
  gene_order = NULL,
  sample.by = NULL,
  allele_counts = NULL,
  reference_counts = NULL,
  output_dir = NULL,
  prefix = "CNV",
  tool_name = "CNV",
  store_matrix = TRUE,
  verbose = TRUE,
  srt = NULL,
  ...
)
Source
The example uses expression and cell annotations from GSE70630.
Arguments
- object
A
Seuratobject.- method
CNA/CNV method. Supported methods are
"copykat","fastCNV","scevan","infercnv", and"numbat".- assay
Assay to use.
NULLuses the default assay.- layer
Assay layer used as the expression matrix.
- group.by
Optional metadata column forwarded to supported methods and stored as cell annotation.
- reference.by
Metadata column identifying reference/normal cells. Required for
"infercnv"and"fastCNV".- reference
Reference labels in
reference.by.- genome
Reference genome label.
- gene_order
Gene coordinate table or a path to one. The table should contain gene, chromosome, start, and end columns. If
NULL, SCOP tries to resolve these columns from assay feature metadata.- sample.by
Optional sample metadata column.
- allele_counts
Allele count table for
"numbat". This is forwarded tonumbat::run_numbat()asdf_allele.- reference_counts
Reference expression profile for
"numbat". This is forwarded tonumbat::run_numbat()aslambdas_ref.- output_dir
Optional output directory.
- prefix
Prefix for metadata columns.
- tool_name
Name used for
srt@tools.- store_matrix
Whether to store the normalized CNV matrix in
srt@tools[[tool_name]].- verbose
Whether to print the message. Default is
TRUE.- srt
Deprecated alias for
object; supply exactly one of the two. It will be removed in scop 1.0.0.- ...
Arguments passed to other functions.
Examples
base_url <- paste0(
  "https://raw.githubusercontent.com/broadinstitute/infercnv/",
  "1def6c930335d00ddf251474abb0a31a1c74b0da/inst/extdata/"
)
files <- c("oligodendroglioma_expression_downsampled.counts.matrix.gz",
           "oligodendroglioma_annotations_downsampled.txt")
counts_file <- tempfile(fileext = ".gz")
labels_file <- tempfile()
utils::download.file(paste0(base_url, files[1]), counts_file, mode = "wb")
utils::download.file(paste0(base_url, files[2]), labels_file, mode = "wb")
counts <- as.matrix(utils::read.delim(counts_file, row.names = 1, check.names = FALSE))
labels <- utils::read.delim(labels_file, header = FALSE, row.names = 1)
cohort <- labels[colnames(counts), 1]
colnames(counts) <- paste0("cell_", colnames(counts))
srt <- SeuratObject::CreateSeuratObject(Matrix::Matrix(counts, sparse = TRUE))
srt$group <- factor(
  ifelse(startsWith(cohort, "malignant_"), sub("^malignant_", "Tumor ", cohort), "Reference"),
  levels = c("Reference", "Tumor 93", "Tumor 97", "Tumor MGH36", "Tumor MGH53")
)
set.seed(42)
srt <- RunCNV(srt, method = "copykat", reference.by = "group", reference = "Reference")
#> ℹ [2026-11-12 05:25:46] Run copykat CNV backend with 10338 features and 184 cells
#> ℹ [2026-11-12 05:25:46] Attached copykat to the search path so its lazy data objects are resolvable by the backend
#> [1] "running copykat v1.2.5"
#> [1] "step1: read and filter data ..."
#> [1] "10338 genes, 184 cells in raw data"
#> [1] "9450 genes past LOW.DR filtering"
#> [1] "step 2: annotations gene coordinates ..."
#> [1] "start annotation ..."
#> [1] "step 3: smoothing data with dlm ..."
#> [1] "step 4: measuring baselines ..."
#> [1] "42 known normal cells found in dataset"
#> [1] "run with known normal..."
#> [1] "baseline is from known input"
#> [1] "step 5: segmentation..."
#> [1] "step 6: convert to genomic bins..."
#> [1] "step 7: re-adjust baseline ..."
#> [1] "step 8: final prediction ..."
#> [1] "step 9: saving results..."
#> [1] "step 10: ploting heatmap ..."
#> Time difference of 1.171434 mins
#> ✔ [2026-11-12 05:26:56] copykat CNV results stored in `srt@tools[[CNV]]`
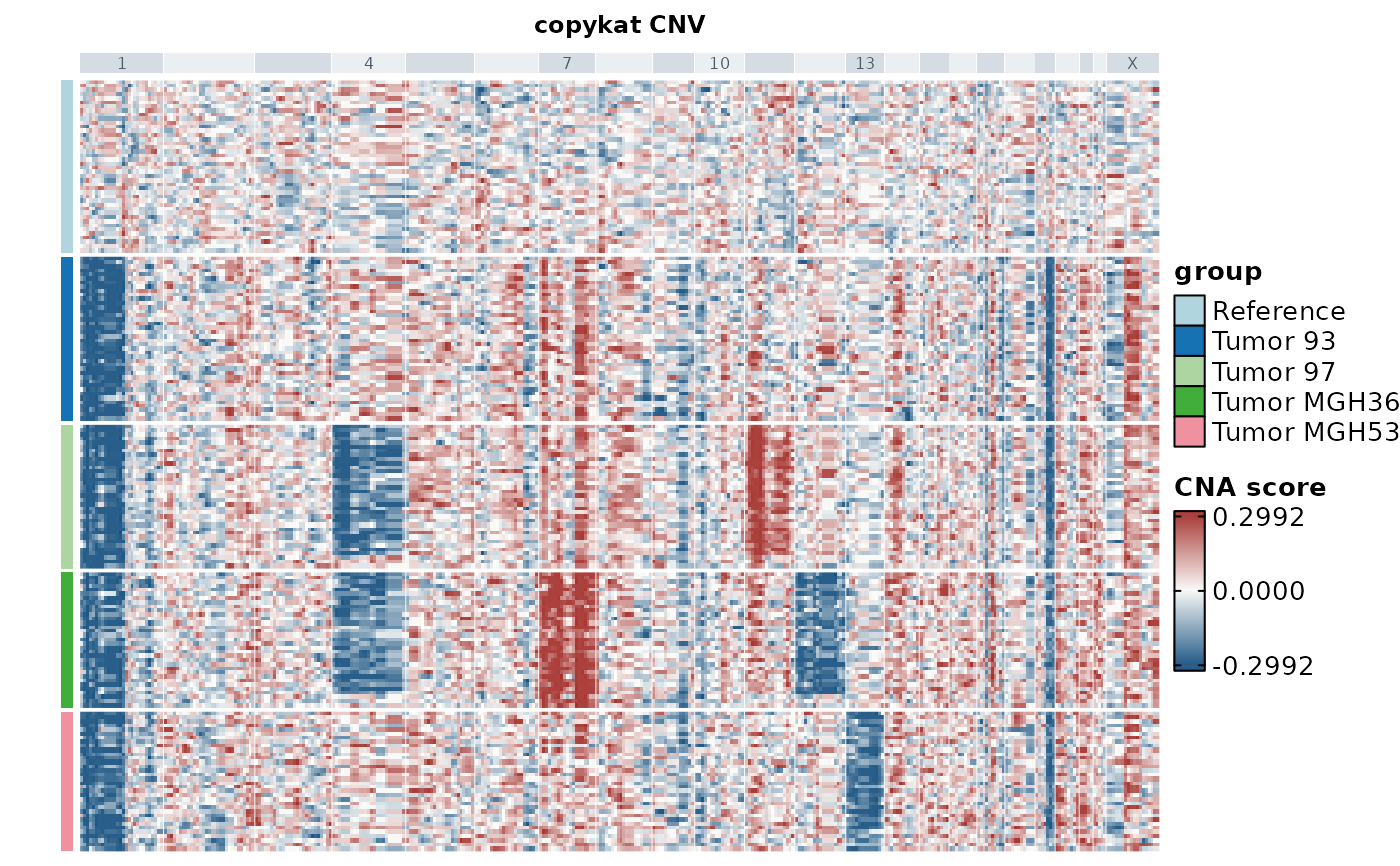
CNVPlot(srt, group.by = "group", heatmap_limits = "auto")