Annotate single cells using scmap
Usage
RunScmap(
  srt_query,
  srt_ref,
  ref_group = NULL,
  query_assay = "RNA",
  ref_assay = "RNA",
  method = "scmapCluster",
  nfeatures = 500,
  threshold = 0.5,
  k = 10,
  verbose = TRUE
)
Arguments
- srt_query
An object of class Seurat to be annotated with cell types.
- srt_ref
An object of class Seurat storing the reference cells.
- ref_group
Column name in the
srt_refmetadata that represents the cell grouping.- query_assay
Assay to be used for the query data. Default is the default assay of the
srt_queryobject.- ref_assay
Assay to be used for the reference data. Default is the default assay of the
srt_refobject.- method
The method to be used for scmap analysis. Can be any of
"scmapCluster"or"scmapCell".- nfeatures
The number of top features to be selected.
- threshold
The threshold value on similarity to determine if a cell is assigned to a cluster. This should be a value between
0and1.- k
Number of clusters per group for k-means clustering when
methodis"scmapCell".- verbose
Whether to print the message. Default is
TRUE.
Examples
data(panc8_sub)
keep <- panc8_sub$celltype %in% c("ductal", "alpha", "beta")
panc8_sub <- Seurat::NormalizeData(panc8_sub[, keep], verbose = FALSE)
reference <- panc8_sub[, panc8_sub$tech != "fluidigmc1"]
query <- panc8_sub[, panc8_sub$tech == "fluidigmc1"]
query <- RunScmap(
  srt_query = query, srt_ref = reference,
  ref_group = "celltype", method = "scmapCluster", nfeatures = 200,
  verbose = FALSE
)
#> ℹ [2026-10-11 23:47:45] Data type is log-normalized
#> ℹ [2026-10-11 23:47:46] Data type is log-normalized
query <- RunStandardWorkflow(query, verbose = FALSE, linear_reduction_dims = 10)
#> ℹ [2026-10-11 23:47:47] Skip `log1p()` because `layer = data` is not "counts"
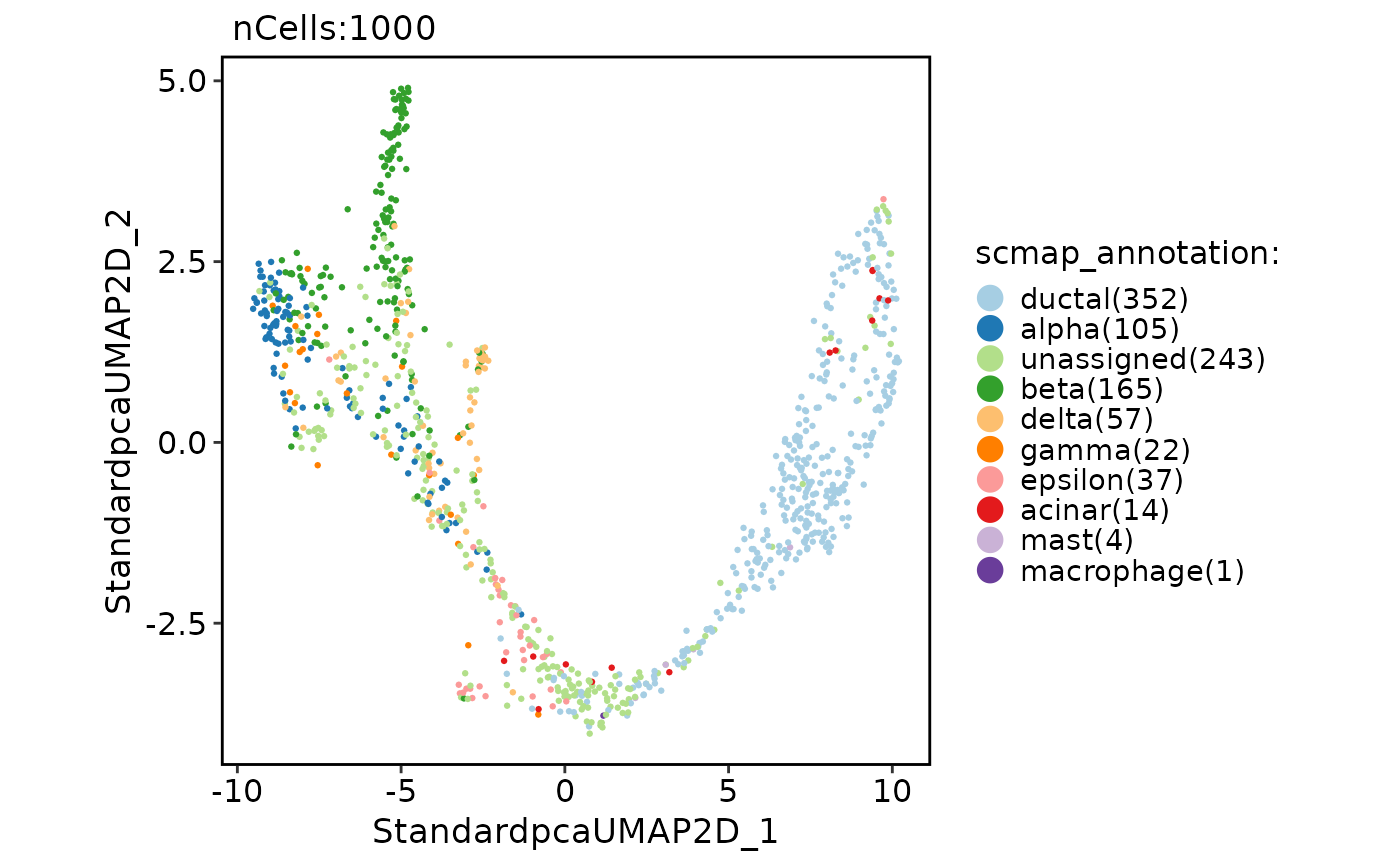
CellDimPlot(
  query,
  group.by = "scmap_annotation",
  label = TRUE, legend.position = "bottom"
)