Ambient RNA decontamination with decontX
Usage
RunDecontX(
  object,
  assay = "RNA",
  group.by = NULL,
  batch = NULL,
  background = NULL,
  background_assay = NULL,
  bg_batch = NULL,
  assay_name = "decontXcounts",
  store_assay = TRUE,
  round_counts = FALSE,
  data_type = NULL,
  seed = 11,
  ...,
  verbose = TRUE,
  srt = NULL
)
Arguments
- object
A
Seuratobject.- assay
Assay to decontaminate.
- group.by, batch
Cell cluster and batch labels passed to
decontX::decontX(). Column name, cell-aligned vector, orNULL.- background
Background / empty-droplet input: a
Seuratobject,SingleCellExperiment, or count matrix.- background_assay
Assay used when
backgroundis aSeuratorSingleCellExperiment.NULLusesassay(Seurat) or"counts"(SCE).- bg_batch
Batch labels for
background.- assay_name, store_assay, round_counts
Store rounded decontaminated counts as a new assay.
- data_type
Optional
CheckDataType()result, used internally to avoid rescanning the count matrix.- seed
Random seed.
- ...
Passed to
decontX::decontX().- verbose
Whether to print messages.
- srt
Deprecated alias for
object; supply exactly one of the two. It will be removed in scop 1.0.0.
Value
A Seurat object with decontX contamination in meta.data and
optional decontaminated counts in a new assay.
Examples
data(pancreas_sub)
pancreas_sub <- RunStandardWorkflow(pancreas_sub)
#> ℹ [2026-10-11 23:14:05] Start standard processing workflow...
#> ℹ [2026-10-11 23:14:05] Checking a list of <Seurat>...
#> ! [2026-10-11 23:14:05] Data 1/1 of the `srt_list` is "unknown"
#> ℹ [2026-10-11 23:14:05] Perform `NormalizeData()` with `normalization.method = 'LogNormalize'` on 1/1 of `srt_list`...
#> ℹ [2026-10-11 23:14:05] Perform `FindVariableFeatures()` on 1/1 of `srt_list`...
#> ℹ [2026-10-11 23:14:05] Use the separate HVF from `srt_list`
#> ℹ [2026-10-11 23:14:05] Number of available HVF: 2000
#> ℹ [2026-10-11 23:14:05] Finished check
#> ℹ [2026-10-11 23:14:05] Perform `ScaleData()`
#> ℹ [2026-10-11 23:14:05] Perform pca linear dimension reduction
#> ℹ [2026-10-11 23:14:06] Use stored estimated dimensions 1:23 for Standardpca
#> ℹ [2026-10-11 23:14:06] Perform `Seurat::FindClusters()` with `cluster_algorithm = 'louvain'` and `cluster_resolution = 0.6`
#> ℹ [2026-10-11 23:14:06] Reorder clusters...
#> ℹ [2026-10-11 23:14:06] Skip `log1p()` because `layer = data` is not "counts"
#> ℹ [2026-10-11 23:14:06] Perform umap nonlinear dimension reduction
#> ✔ [2026-10-11 23:14:13] Standard processing workflow completed
pancreas_sub <- RunDecontX(
pancreas_sub,
group.by = "CellType"
)
#> ℹ [2026-10-11 23:14:13] Running decontX
#> ℹ [2026-10-11 23:14:13] Data type is raw counts
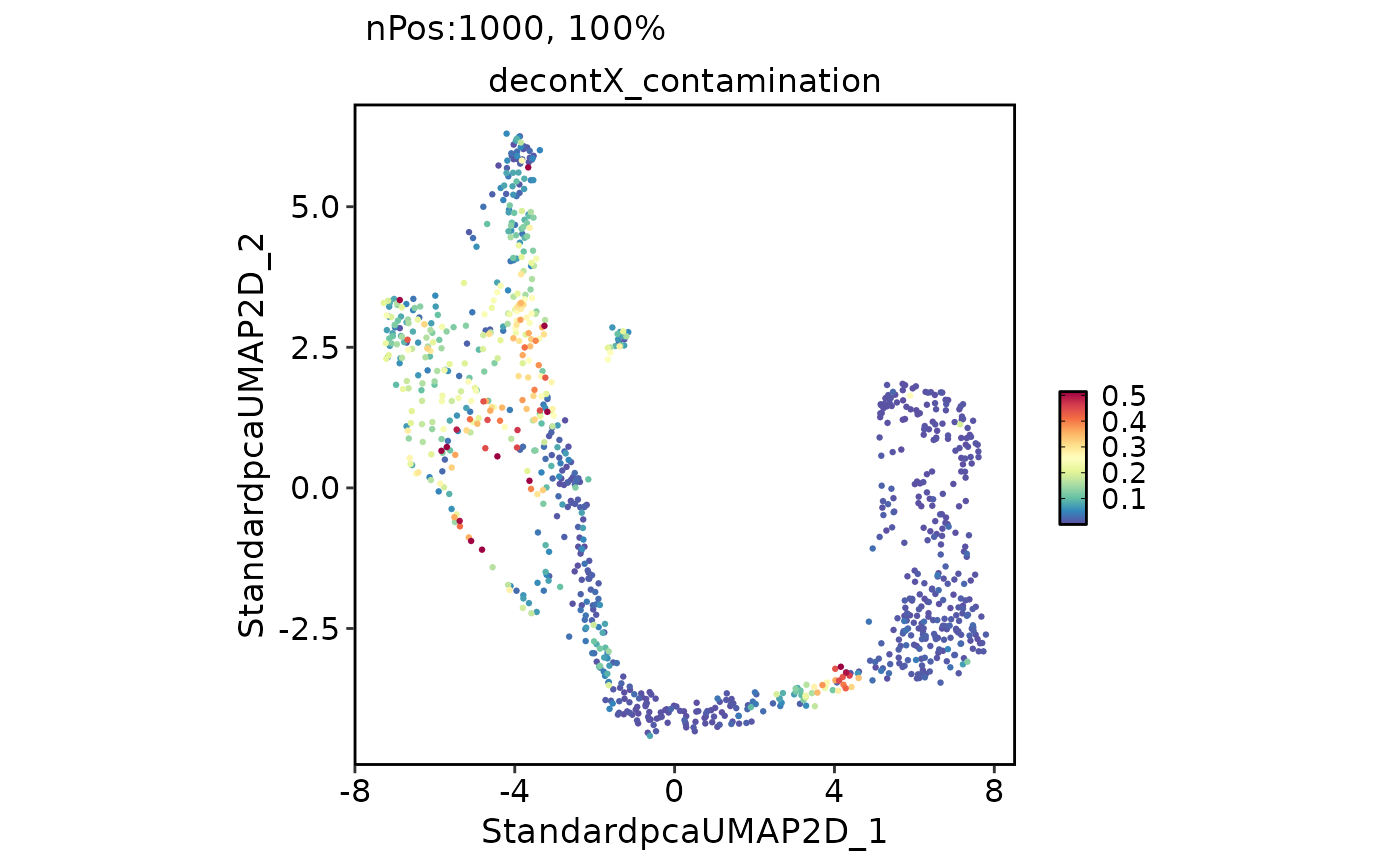
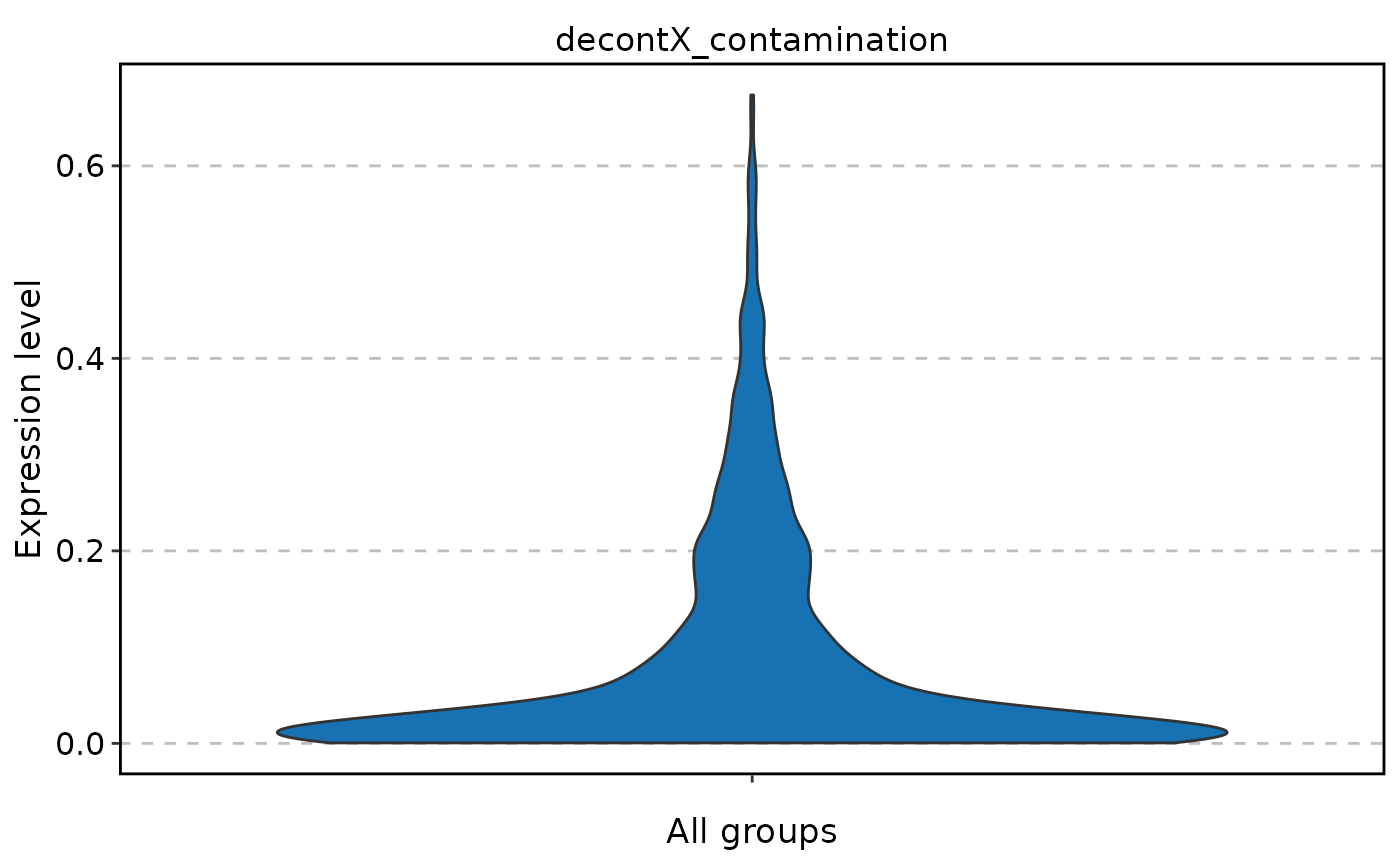
#> ℹ [2026-10-11 23:14:29] decontX contamination (median/mean/max): 0.0272 / 0.0875 / 0.6737
#> ℹ [2026-10-11 23:14:29] decontX assay stored as decontXcounts
#> ✔ [2026-10-11 23:14:29] decontX decontamination completed
FeatureStatPlot(
pancreas_sub,
stat.by = "decontX_contamination"
)
 FeatureDimPlot(
pancreas_sub,
features = "decontX_contamination"
)